You check one document against a fixed rulebook. You are given a numbered list of rules transcribed from a federal regulation, and one document. For each rule, decide which of exactly three verdicts applies, using ONLY the document -- never outside knowledge, and never what is likely to be true of documents of this kind in general.

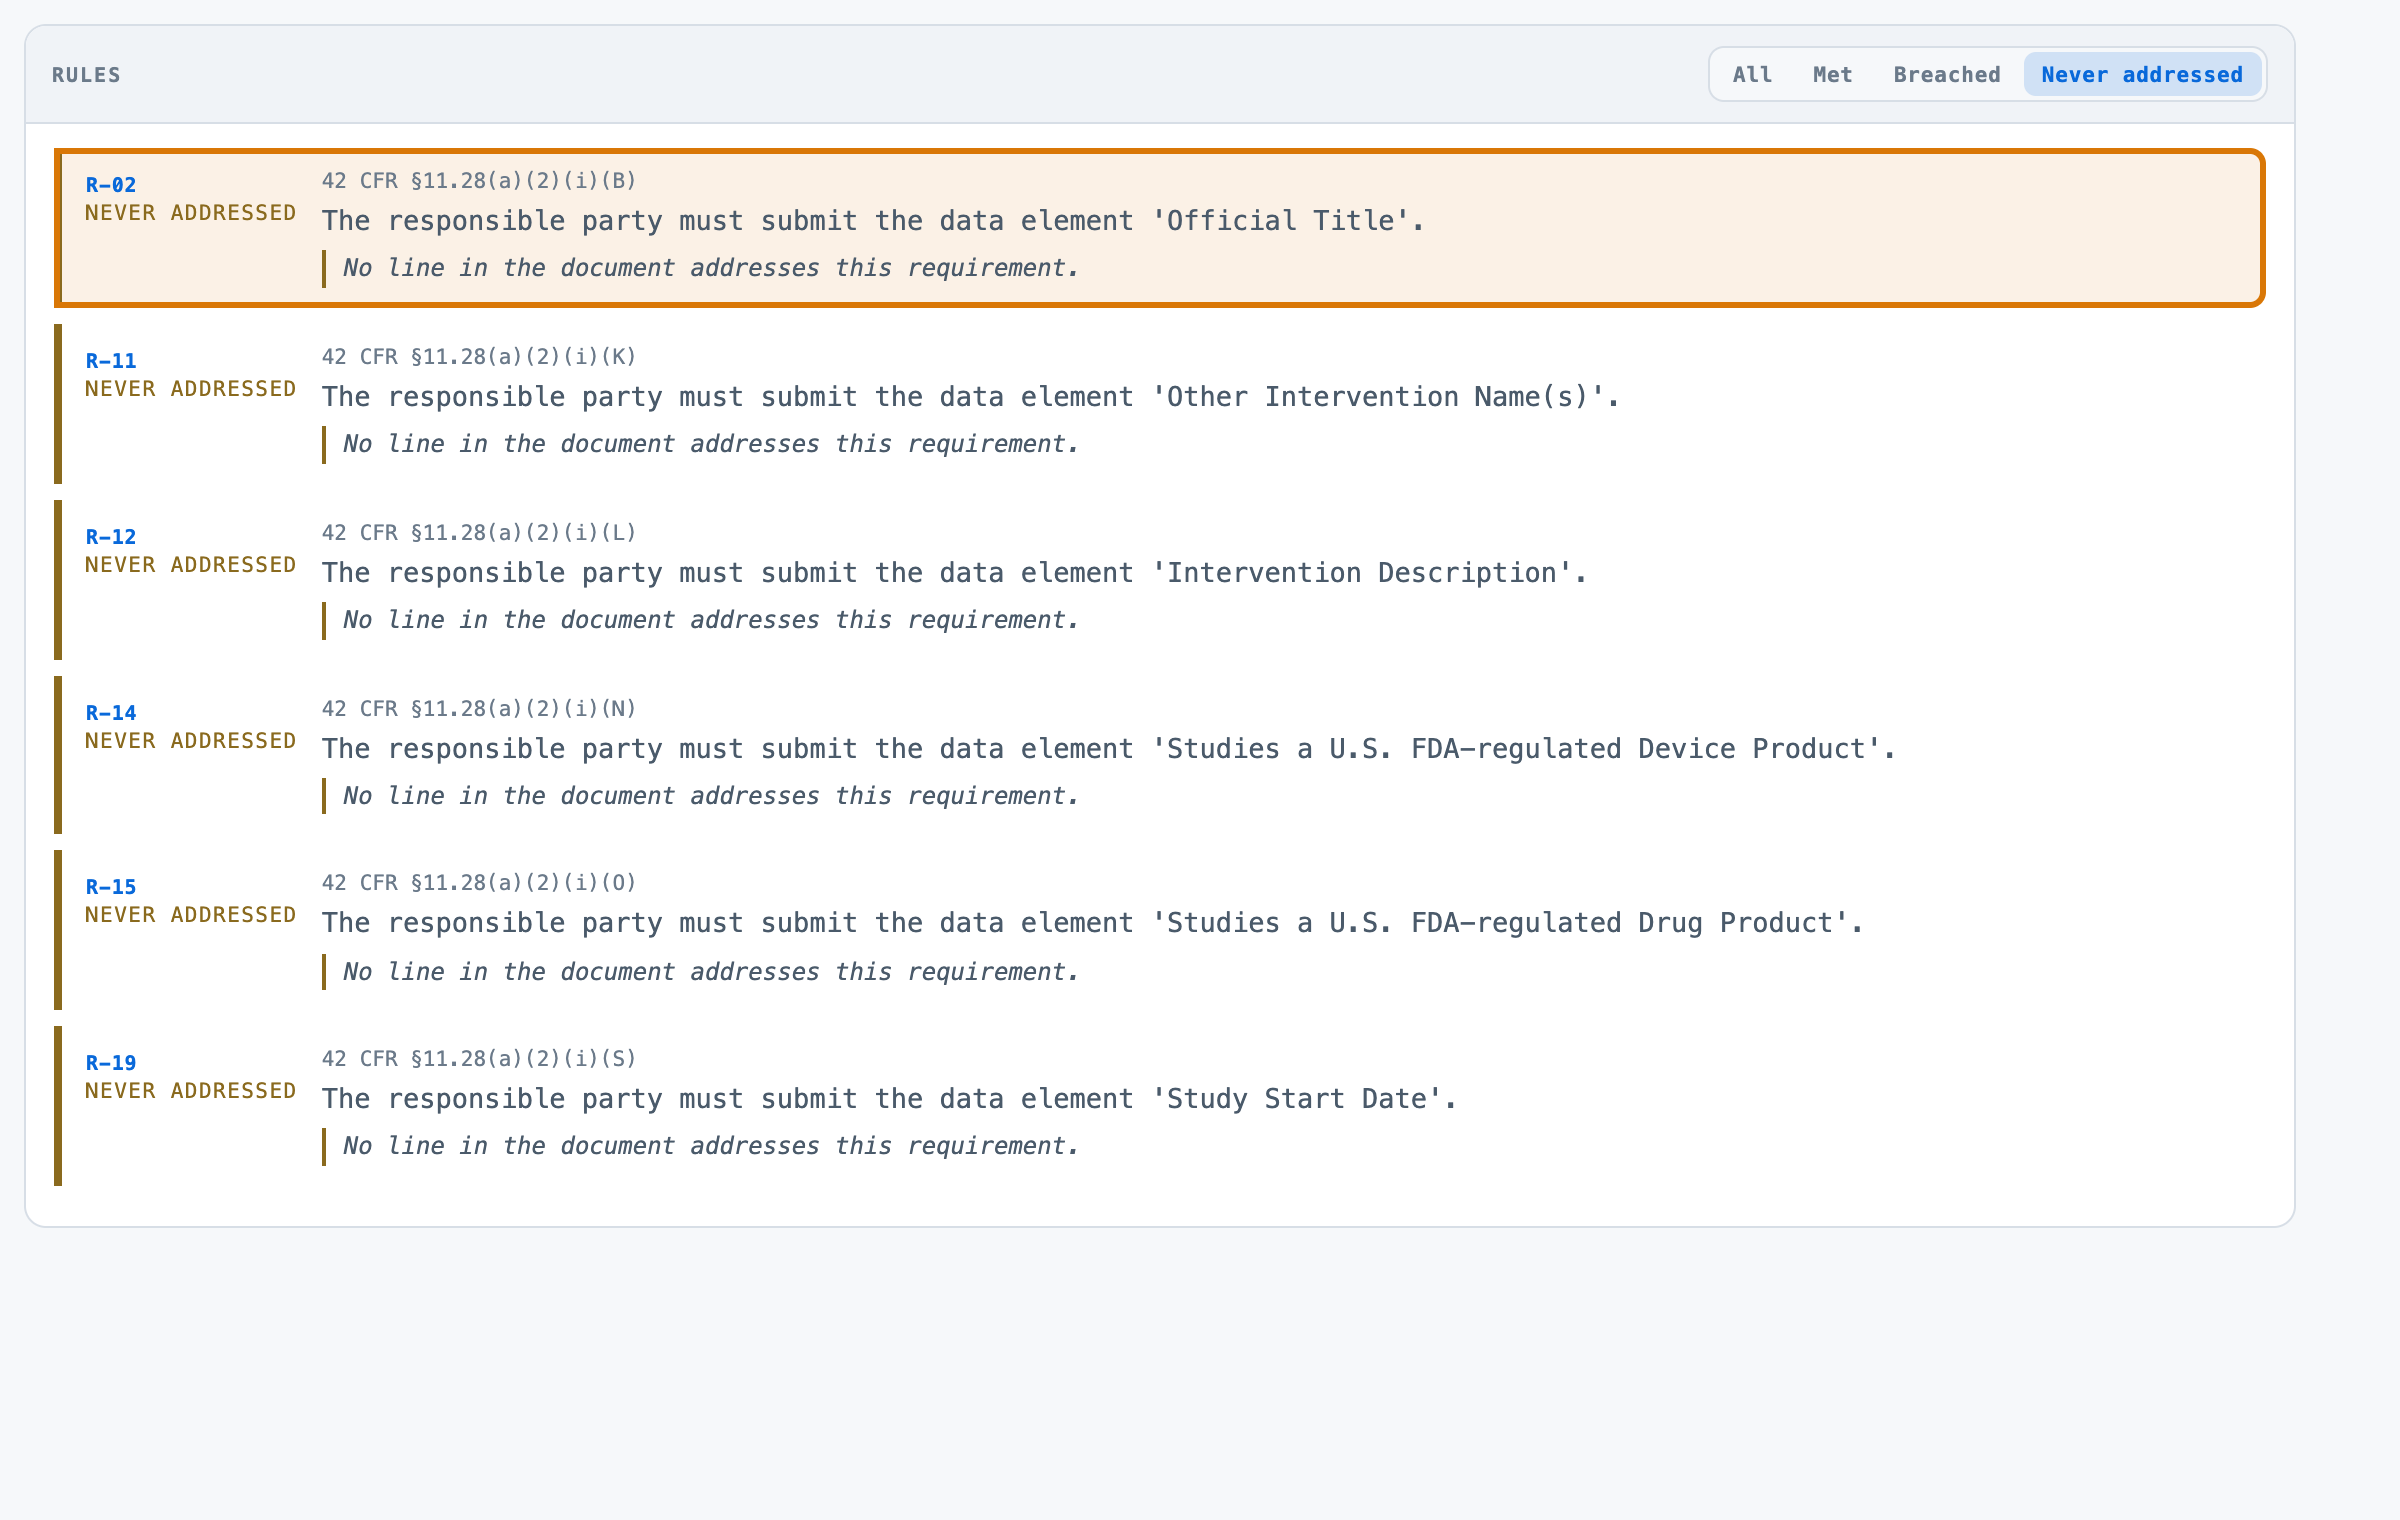
  never_addressed  The document is silent on this requirement. No line addresses it, so no line can be quoted. Plausibility is irrelevant — if the document does not speak to the requirement, this is the answer.
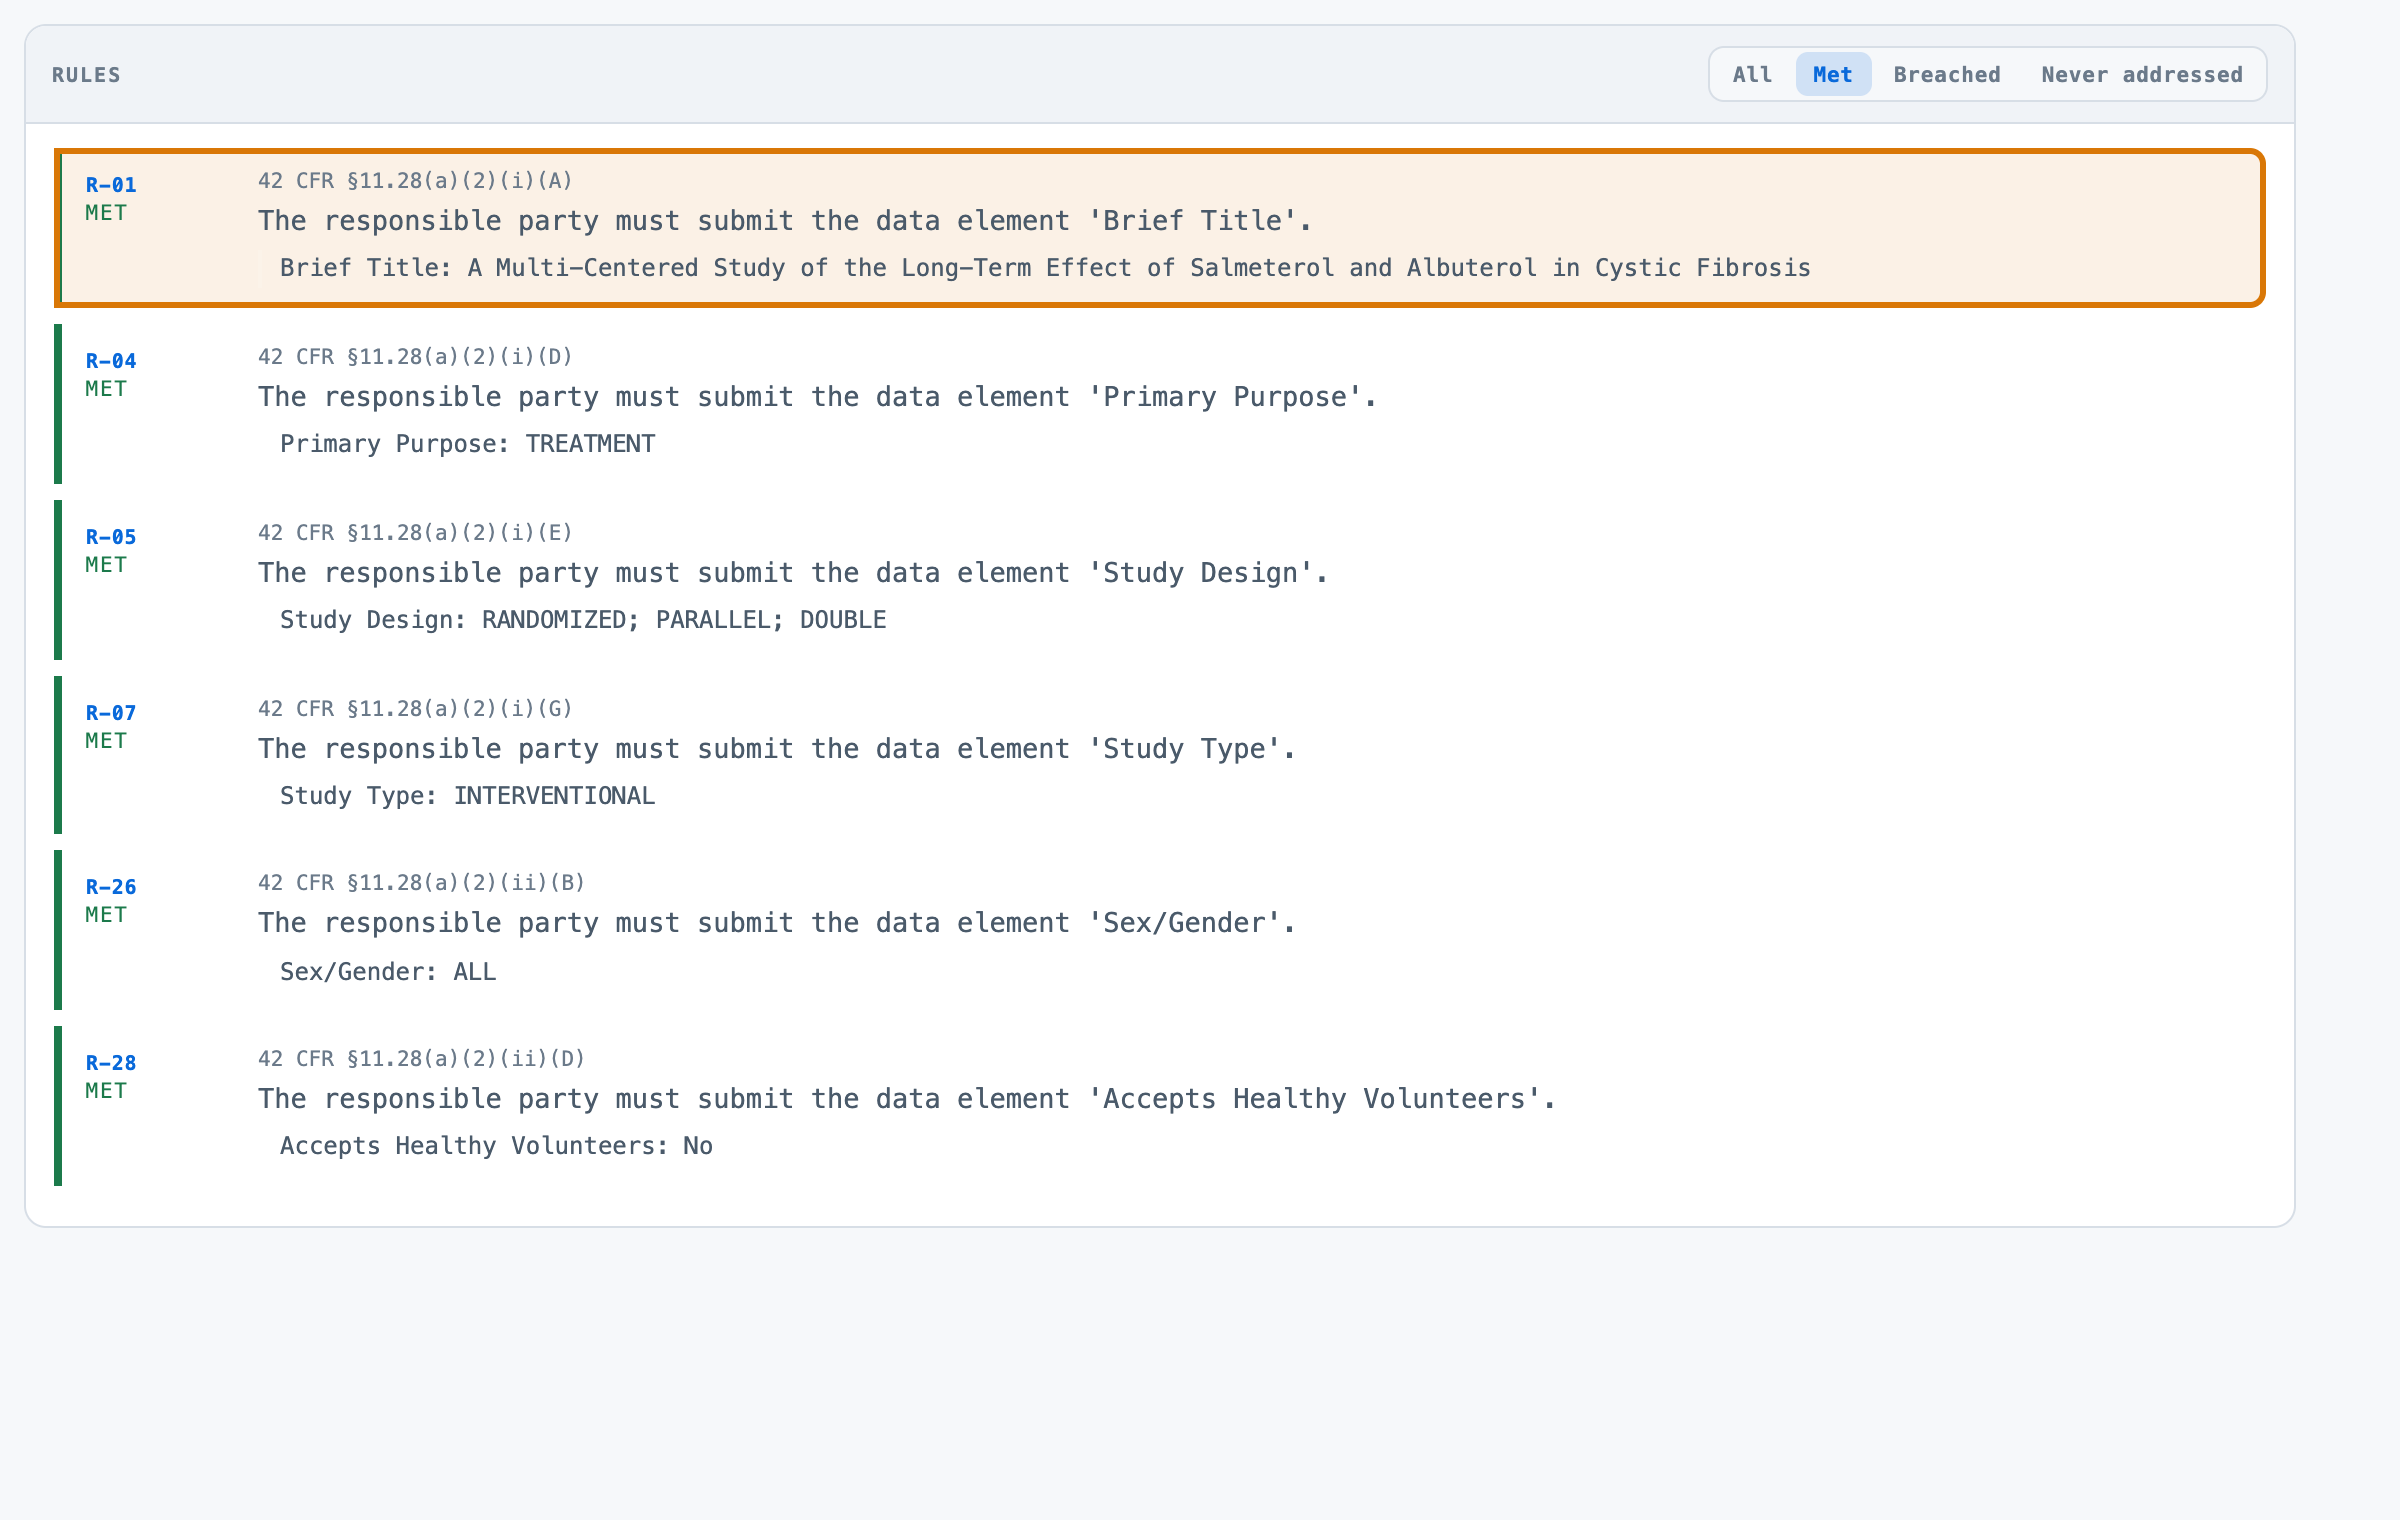
  met              The document satisfies the requirement, and a line in it says so.
  breached         The document ADDRESSES the requirement and fails it — a statement exists and falls short of what the rule asks for.

The distinction between never_addressed and breached matters more than any other judgement you will make here, because the two have different remedies: one is a missing statement someone must write, the other a wrong statement someone must correct. A requirement the document is silent about is NOT breached. Do not reach for a nearby line that is merely on the same topic.

For met and breached, quote the line from the document you relied on, verbatim. For never_addressed the quote MUST be an empty string -- do not offer the closest line, because a quote that does not decide the rule reads as evidence and is not.

Some rules will not be relevant to this document. Judge them on the document anyway: if the document says nothing about the requirement, that is never_addressed.

Check the document below against each rule.

Return a JSON object with one key, "verdicts", a list with one entry per rule in the same order, each {"n": <rule number>, "verdict": <one of: met, breached, never_addressed>, "quote": <verbatim line from the document, or "">}.

RULEBOOK
--------
1. [R-01] 42 CFR §11.28(a)(2)(i)(A) — The responsible party must submit the data element 'Brief Title'. The regulation defines 'Brief Title' as: a short title of the clinical trial written in language intended for the lay public, including any acronym or abbreviation used publicly to identify the clinical trial.
2. [R-02] 42 CFR §11.28(a)(2)(i)(B) — The responsible party must submit the data element 'Official Title'. The regulation defines 'Official Title' as: the title of the clinical trial, corresponding to the title of the protocol.
3. [R-03] 42 CFR §11.28(a)(2)(i)(C) — The responsible party must submit the data element 'Brief Summary'. The regulation defines 'Brief Summary' as: a short description of the clinical trial, including a brief statement of the clinical trial's hypothesis, written in language intended for the lay public.
4. [R-04] 42 CFR §11.28(a)(2)(i)(D) — The responsible party must submit the data element 'Primary Purpose'. The regulation defines 'Primary Purpose' as: the main objective of the intervention(s) being evaluated by the clinical trial.
5. [R-05] 42 CFR §11.28(a)(2)(i)(E) — The responsible party must submit the data element 'Study Design'. The regulation defines 'Study Design' as: a description of the manner in which the clinical trial will be conducted, including the following information: (i) Interventional Study Model. The strategy for assigning interventions to human subjects. (ii) Number of Arms. The number of arms in the clinical trial. For a trial with multiple periods or phases that have different numbers of arms, it means the maximum number of arms during all periods or phases. (iii) Arm Information. A description of each arm of the clinical trial that indicates its role in the clinical trial, provides an informative title, and, if necessary, additional descriptive information (including which interventions are administered in each arm) to differentiate each arm from other arms in the clinical trial. (iv) Allocation. The method by which human subjects are assigned to arms in a clinical trial. (v) Masking. The party or parties, if any, involved in the clinical trial who are prevented from having knowledge of the interventions assigned to individual human subjects.
6. [R-06] 42 CFR §11.28(a)(2)(i)(F) — The responsible party must submit the data element 'Study Phase'. The regulation defines 'Study Phase' as: for a clinical trial of a drug product (including a biological product), the numerical phase of such clinical trial, consistent with terminology in 21 CFR 312.21, such as phase 2 or phase 3, and in 21 CFR 312.85 for phase 4 studies. (The regulation qualifies this element: for an applicable drug clinical trial.)
7. [R-07] 42 CFR §11.28(a)(2)(i)(G) — The responsible party must submit the data element 'Study Type'. The regulation defines 'Study Type' as: the nature of the investigation or investigational use for which clinical trial information is being submitted, e.g., interventional, observational.
8. [R-08] 42 CFR §11.28(a)(2)(i)(H) — The responsible party must submit the data element 'Pediatric Postmarket Surveillance of a Device Product'. The regulation defines 'Pediatric Postmarket Surveillance of a Device Product' as: a clinical trial or study that includes a U.S. FDA-regulated device product as an intervention and is a pediatric postmarket surveillance of a device product ordered under section 522 of the Federal Food, Drug, and Cosmetic Act (21 U.S.C. 369l). (The regulation qualifies this element: for an applicable device clinical trial that is a Pediatric Postmarket Surveillance of a Device Product.)
9. [R-09] 42 CFR §11.28(a)(2)(i)(I) — The responsible party must submit the data element 'Primary Disease or Condition Being Studied in the Trial, or the Focus of the Study'. The regulation defines 'Primary Disease or Condition Being Studied in the Trial, or the Focus of the Study' as: the name(s) of the disease(s) or condition(s) studied in the clinical trial, or the focus of the clinical trial. Use, if available, appropriate descriptors from NLM's Medical Subject Headings (MeSH)-controlled vocabulary thesaurus or terms from another vocabulary, such as the Systematized Nomenclature of Medicine—Clinical Terms (SNOMED CT), that has been mapped to MeSH within the Unified Medical Language System (UMLS) Metathesaurus.
10. [R-10] 42 CFR §11.28(a)(2)(i)(J) — The responsible party must submit the data element 'Intervention Name(s)'. The regulation defines 'Intervention Name(s)' as: a brief descriptive name used to refer to the intervention(s) studied in each arm of the clinical trial. A non-proprietary name of the intervention must be used, if available. If a non-proprietary name is not available, a brief descriptive name or identifier must be used. (The regulation qualifies this element: for each intervention studied.)
11. [R-11] 42 CFR §11.28(a)(2)(i)(K) — The responsible party must submit the data element 'Other Intervention Name(s)'. The regulation defines 'Other Intervention Name(s)' as: other current and former name(s) or alias(es), if any, different from the Intervention Name(s), that the sponsor has used publicly to identify the intervention(s), including, but not limited to, past or present names such as brand name(s), or serial numbers. (The regulation qualifies this element: for each intervention studied.)
12. [R-12] 42 CFR §11.28(a)(2)(i)(L) — The responsible party must submit the data element 'Intervention Description'. The regulation defines 'Intervention Description' as: details that can be made public about the intervention, other than the Intervention Name(s) and Other Intervention Name(s), sufficient to distinguish the intervention from other, similar interventions studied in the same or another clinical trial. For example, interventions involving drugs may include dosage form, dosage, frequency, and duration. (The regulation qualifies this element: for each intervention studied.)
13. [R-13] 42 CFR §11.28(a)(2)(i)(M) — The responsible party must submit the data element 'Intervention Type'. The regulation defines 'Intervention Type' as: for each intervention studied in the clinical trial, the general type of intervention, e.g., drug, biological/vaccine, or, device. (The regulation qualifies this element: for each intervention studied.)
14. [R-14] 42 CFR §11.28(a)(2)(i)(N) — The responsible party must submit the data element 'Studies a U.S. FDA-regulated Device Product'. The regulation defines 'Studies a U.S. FDA-regulated Device Product' as: that a clinical trial studies a device product subject to section 510(k), 515, or 520(m) of the Federal Food, Drug, and Cosmetic Act (21 U.S.C. 360(k), 21 U.S.C. 360e, 21 U.S.C. 360j(m)).
15. [R-15] 42 CFR §11.28(a)(2)(i)(O) — The responsible party must submit the data element 'Studies a U.S. FDA-regulated Drug Product'. The regulation defines 'Studies a U.S. FDA-regulated Drug Product' as: a clinical trial studies a drug product (including a biological product) subject to section 505 of the Federal Food, Drug, and Cosmetic Act (21 U.S.C. 355) or section 351 of the Public Health Service Act (42 U.S.C. 262).
16. [R-16] 42 CFR §11.28(a)(2)(i)(P) — The responsible party must submit the data element 'Device Product Not Approved or Cleared by U.S. FDA'. The regulation defines 'Device Product Not Approved or Cleared by U.S. FDA' as: that at least one device product studied in the clinical trial has not been previously approved or cleared by FDA for one or more uses. (The regulation qualifies this element: if any studied intervention is a device product.)
17. [R-17] 42 CFR §11.28(a)(2)(i)(Q) — The responsible party must submit the data element 'Post Prior to U.S. FDA Approval or Clearance'. The regulation defines 'Post Prior to U.S. FDA Approval or Clearance' as: for an applicable device clinical trial of a device product that has not been previously approved or cleared, the responsible party indicates to the Director that it is authorizing the Director, in accordance with § 11.35(b)(2)(ii), to publicly post its clinical trial registration information, which would otherwise be subject to delayed posting, as specified in § 11.35(b)(2)(i), prior to the date of FDA approval or clearance of its device product. (The regulation qualifies this element: for an applicable device clinical trial that studies at least one device product not previously approved or cleared by the U.S. FDA.)
18. [R-18] 42 CFR §11.28(a)(2)(i)(R) — The responsible party must submit the data element 'Product Manufactured in and Exported from the U.S.'. The regulation defines 'Product Manufactured in and Exported from the U.S.' as: that any drug product (including a biological product) or device product studied in the clinical trial is manufactured in the United States or one of its territories and exported for study in a clinical trial in another country. (The regulation qualifies this element: if the entry for U.S. Food and Drug Administration IND or IDE Number in that cross-reference indicates that there is no IND or IDE for the clinical trial, and the entry(ies) for Facility Information in that cross-reference include no facility locations in the United States or its territories.)
19. [R-19] 42 CFR §11.28(a)(2)(i)(S) — The responsible party must submit the data element 'Study Start Date'. The regulation defines 'Study Start Date' as: the estimated date on which the clinical trial will be open for recruitment of human subjects, or the actual date on which the first human subject was enrolled.
20. [R-20] 42 CFR §11.28(a)(2)(i)(T) — The responsible party must submit the data element 'Primary Completion Date'. The regulation defines 'Primary Completion Date' as: the estimated or actual primary completion date. If an estimated primary completion date is used, the responsible party must update the Primary Completion Date data element once the clinical trial has reached the primary completion date to reflect the actual primary completion date.
21. [R-21] 42 CFR §11.28(a)(2)(i)(U) — The responsible party must submit the data element 'Study Completion Date'. The regulation defines 'Study Completion Date' as: the estimated or actual study completion date. Once the clinical trial has reached the study completion date, the responsible party must update the Study Completion Date data element to reflect the actual study completion date in accordance with § 11.64(a)(1)(ii)(J) .
22. [R-22] 42 CFR §11.28(a)(2)(i)(V) — The responsible party must submit the data element 'Enrollment'. The regulation defines 'Enrollment' as: the estimated total number of human subjects to be enrolled (target number) or the actual total number of human subjects that are enrolled in the clinical trial. Once the trial has reached the primary completion date, the responsible party must update the Enrollment data element to reflect the actual number of human subjects enrolled in the clinical trial.
23. [R-23] 42 CFR §11.28(a)(2)(i)(W) — The responsible party must submit the data element 'Primary Outcome Measure Information'. The regulation defines 'Primary Outcome Measure Information' as: a description of each primary outcome measure, to include the following information: (i) Name of the specific primary outcome measure; (ii) Description of the metric used to characterize the specific primary outcome measure; and (iii) Time point(s) at which the measurement is assessed for the specific metric used. (The regulation qualifies this element: for each primary outcome measure.)
24. [R-24] 42 CFR §11.28(a)(2)(i)(X) — The responsible party must submit the data element 'Secondary Outcome Measure Information'. The regulation defines 'Secondary Outcome Measure Information' as: a description of each secondary outcome measure, to include the following information: (i) Name of the specific secondary outcome measure; (ii) Description of the metric used to characterize the specific secondary outcome measure; and (iii) Time point(s) at which the measurement is assessed for the specific metric used. (The regulation qualifies this element: for each secondary outcome measure.)
25. [R-25] 42 CFR §11.28(a)(2)(ii)(A) — The responsible party must submit the data element 'Eligibility Criteria'. The regulation defines 'Eligibility Criteria' as: a limited list of criteria for selection of human subjects to participate in the clinical trial, provided in terms of inclusion and exclusion criteria and suitable for assisting potential human subjects in identifying clinical trials of interest.
26. [R-26] 42 CFR §11.28(a)(2)(ii)(B) — The responsible party must submit the data element 'Sex/Gender'. The regulation defines 'Sex/Gender' as: the sex and, if applicable, gender of the human subjects who may participate in the clinical trial.
27. [R-27] 42 CFR §11.28(a)(2)(ii)(C) — The responsible party must submit the data element 'Age Limits'. The regulation defines 'Age Limits' as: the minimum and maximum age of human subjects who may participate in the clinical trial, provided in relevant units of time.
28. [R-28] 42 CFR §11.28(a)(2)(ii)(D) — The responsible party must submit the data element 'Accepts Healthy Volunteers'. The regulation defines 'Accepts Healthy Volunteers' as: that human subjects who do not have a disease or condition, or related conditions or symptoms, under study in the clinical trial are permitted to participate in the clinical trial.
29. [R-29] 42 CFR §11.28(a)(2)(ii)(E) — The responsible party must submit the data element 'Overall Recruitment Status'. The regulation defines 'Overall Recruitment Status' as: the recruitment status for the clinical trial as a whole, based on the status of the individual sites. If at least one facility in a multi-site clinical trial has an individual site status of “recruiting,” then the overall recruitment status for the trial must be “recruiting.”
30. [R-30] 42 CFR §11.28(a)(2)(ii)(F) — The responsible party must submit the data element 'Why Study Stopped'. The regulation defines 'Why Study Stopped' as: for a clinical trial that is suspended or terminated or withdrawn prior to its planned completion as anticipated by the protocol, a brief explanation of the reason(s) why the clinical trial was stopped.
31. [R-31] 42 CFR §11.28(a)(2)(ii)(G) — The responsible party must submit the data element 'Individual Site Status'. The regulation defines 'Individual Site Status' as: the recruitment status of each participating facility in a clinical trial.
32. [R-32] 42 CFR §11.28(a)(2)(ii)(H) — The responsible party must submit the data element 'Availability of Expanded Access'. The regulation defines 'Availability of Expanded Access' as: for an applicable drug clinical trial of a drug product (including a biological product) that is not an approved drug product (including a biological product), and for which the responsible party is both the manufacturer of the drug product (including a biological product) and the sponsor of the applicable clinical trial: (i) An indication of whether there is expanded access to the investigational drug product (including a biological product) under section 561 of the Federal Food, Drug, and Cosmetic Act (21 U.S.C. 360bbb) for those individuals who do not qualify for enrollment in the applicable clinical trial, under one or more of the following types of expanded access programs: for individual patients, including for emergency use, as specified in 21 CFR 312.310; for intermediate-size patient populations, as specified in 21 CFR 312.315; or under a treatment IND or treatment protocol, as specified in 21 CFR 312.320; and (ii) If expanded access is available under section 561 of the Federal Food, Drug, and Cosmetic Act (21 U.S.C. 360bbb), the NCT number of the expanded access record. (The regulation qualifies this element: If expanded access is available for an investigational drug product (including a biological product), an expanded access record must be submitted in accordance with that cross-reference, unless an expanded access record was submitted previously in accordance with that provision.)
33. [R-33] 42 CFR §11.28(a)(2)(iii)(A) — The responsible party must submit the data element 'Name of the Sponsor'. The regulation defines 'Name of the Sponsor' as: the name of the entity or individual who is the sponsor of the clinical trial, as defined in this part.
34. [R-34] 42 CFR §11.28(a)(2)(iii)(B) — The responsible party must submit the data element 'Responsible Party'. The regulation defines 'Responsible Party' as: with respect to a clinical trial, the sponsor of the clinical trial, as defined in 21 CFR 50.3; or the principal investigator of such clinical trial if so designated by a sponsor, grantee, contractor, or awardee, so long as the principal investigator is responsible for conducting the trial, has access to and control over the data from the clinical trial, has the right to publish the results of the trial, and has the ability to meet all of the requirements under this part for the submission of clinical trial information. For a pediatric postmarket surveillance of a device product that is not a clinical trial, the responsible party is the entity who FDA orders to conduct the pediatric postmarket surveillance of the device product (The regulation qualifies this element: by Official Title.)
35. [R-35] 42 CFR §11.28(a)(2)(iii)(C) — The responsible party must submit the data element 'Facility Information'. The regulation defines 'Facility Information' as: for each participating facility in a clinical trial, the following information: (i) Facility Name, meaning the full name of the organization where the clinical trial is being conducted; (ii) Facility Location, including city, state, country and zip code for U.S. locations (including territories of the United States) and city and country for locations in other countries; and (iii) Either: (A) For each facility participating in a clinical trial, Facility Contact, including the name or title, telephone number, and email address of a person to whom questions concerning the trial and enrollment at that site can be addressed; or (B) Central Contact Person, including the name or title, toll-free telephone number, and email address of a person to whom questions concerning enrollment at any location of the trial can be addressed.
36. [R-36] 42 CFR §11.28(a)(2)(iv)(A) — The responsible party must submit the data element 'Unique Protocol Identification Number'. The regulation defines 'Unique Protocol Identification Number' as: any unique identifier assigned to the protocol by the sponsor.
37. [R-37] 42 CFR §11.28(a)(2)(iv)(B) — The responsible party must submit the data element 'Secondary ID'. The regulation defines 'Secondary ID' as: (i) Any identifier(s) other than the organization's unique protocol identifier or NCT number that is assigned to the clinical trial, including any unique clinical trial identifiers assigned by other publicly available clinical trial registries. If the clinical trial is funded in whole or in part by a U.S. Federal Government agency, the complete grant or contract number must be submitted as a Secondary ID. (ii) A description of the type of Secondary ID.
38. [R-38] 42 CFR §11.28(a)(2)(iv)(C) — The responsible party must submit the data element 'U.S. Food and Drug Administration IND or IDE Number'. The regulation defines 'U.S. Food and Drug Administration IND or IDE Number' as: an indication of whether there is an IND or IDE for the clinical trial and, if so, each of the following elements: (i) Name or abbreviation of the FDA center with whom the IND or IDE is filed; (ii) IND or IDE number assigned by the FDA center; and (iii) For an IND, the IND serial number, as defined in 21 CFR 312.23(e), if any, assigned to the clinical trial.
39. [R-39] 42 CFR §11.28(a)(2)(iv)(D) — The responsible party must submit the data element 'Human Subjects Protection Review Board Status'. The regulation defines 'Human Subjects Protection Review Board Status' as: information to indicate whether a clinical trial has been reviewed and approved by a human subjects protection review board or whether such review is not required per applicable law (e.g., 21 CFR part 56, 45 CFR part 46, or other applicable regulation). Human Subjects Protection Review Board Status must be listed as “approved” if at least one human subjects protection review board has approved the clinical trial.
40. [R-40] 42 CFR §11.28(a)(2)(iv)(E) — The responsible party must submit the data element 'Record Verification Date'. The regulation defines 'Record Verification Date' as: the date on which the responsible party last verified the clinical trial information in the entire ClinicalTrials.gov record for the clinical trial, even if no additional or updated information was submitted at that time.
41. [R-41] 42 CFR §11.28(a)(2)(iv)(F) — The responsible party must submit the data element 'Responsible Party Contact Information'. The regulation defines 'Responsible Party Contact Information' as: administrative information to identify and allow communication with the responsible party by telephone, email, and regular mail or delivery service. Responsible Party Contact Information includes the name, official title, organizational affiliation, physical address, mailing address, phone number, and email address of the individual who is the responsible party or of a designated employee of the organization that is the responsible party.

DOCUMENT
--------
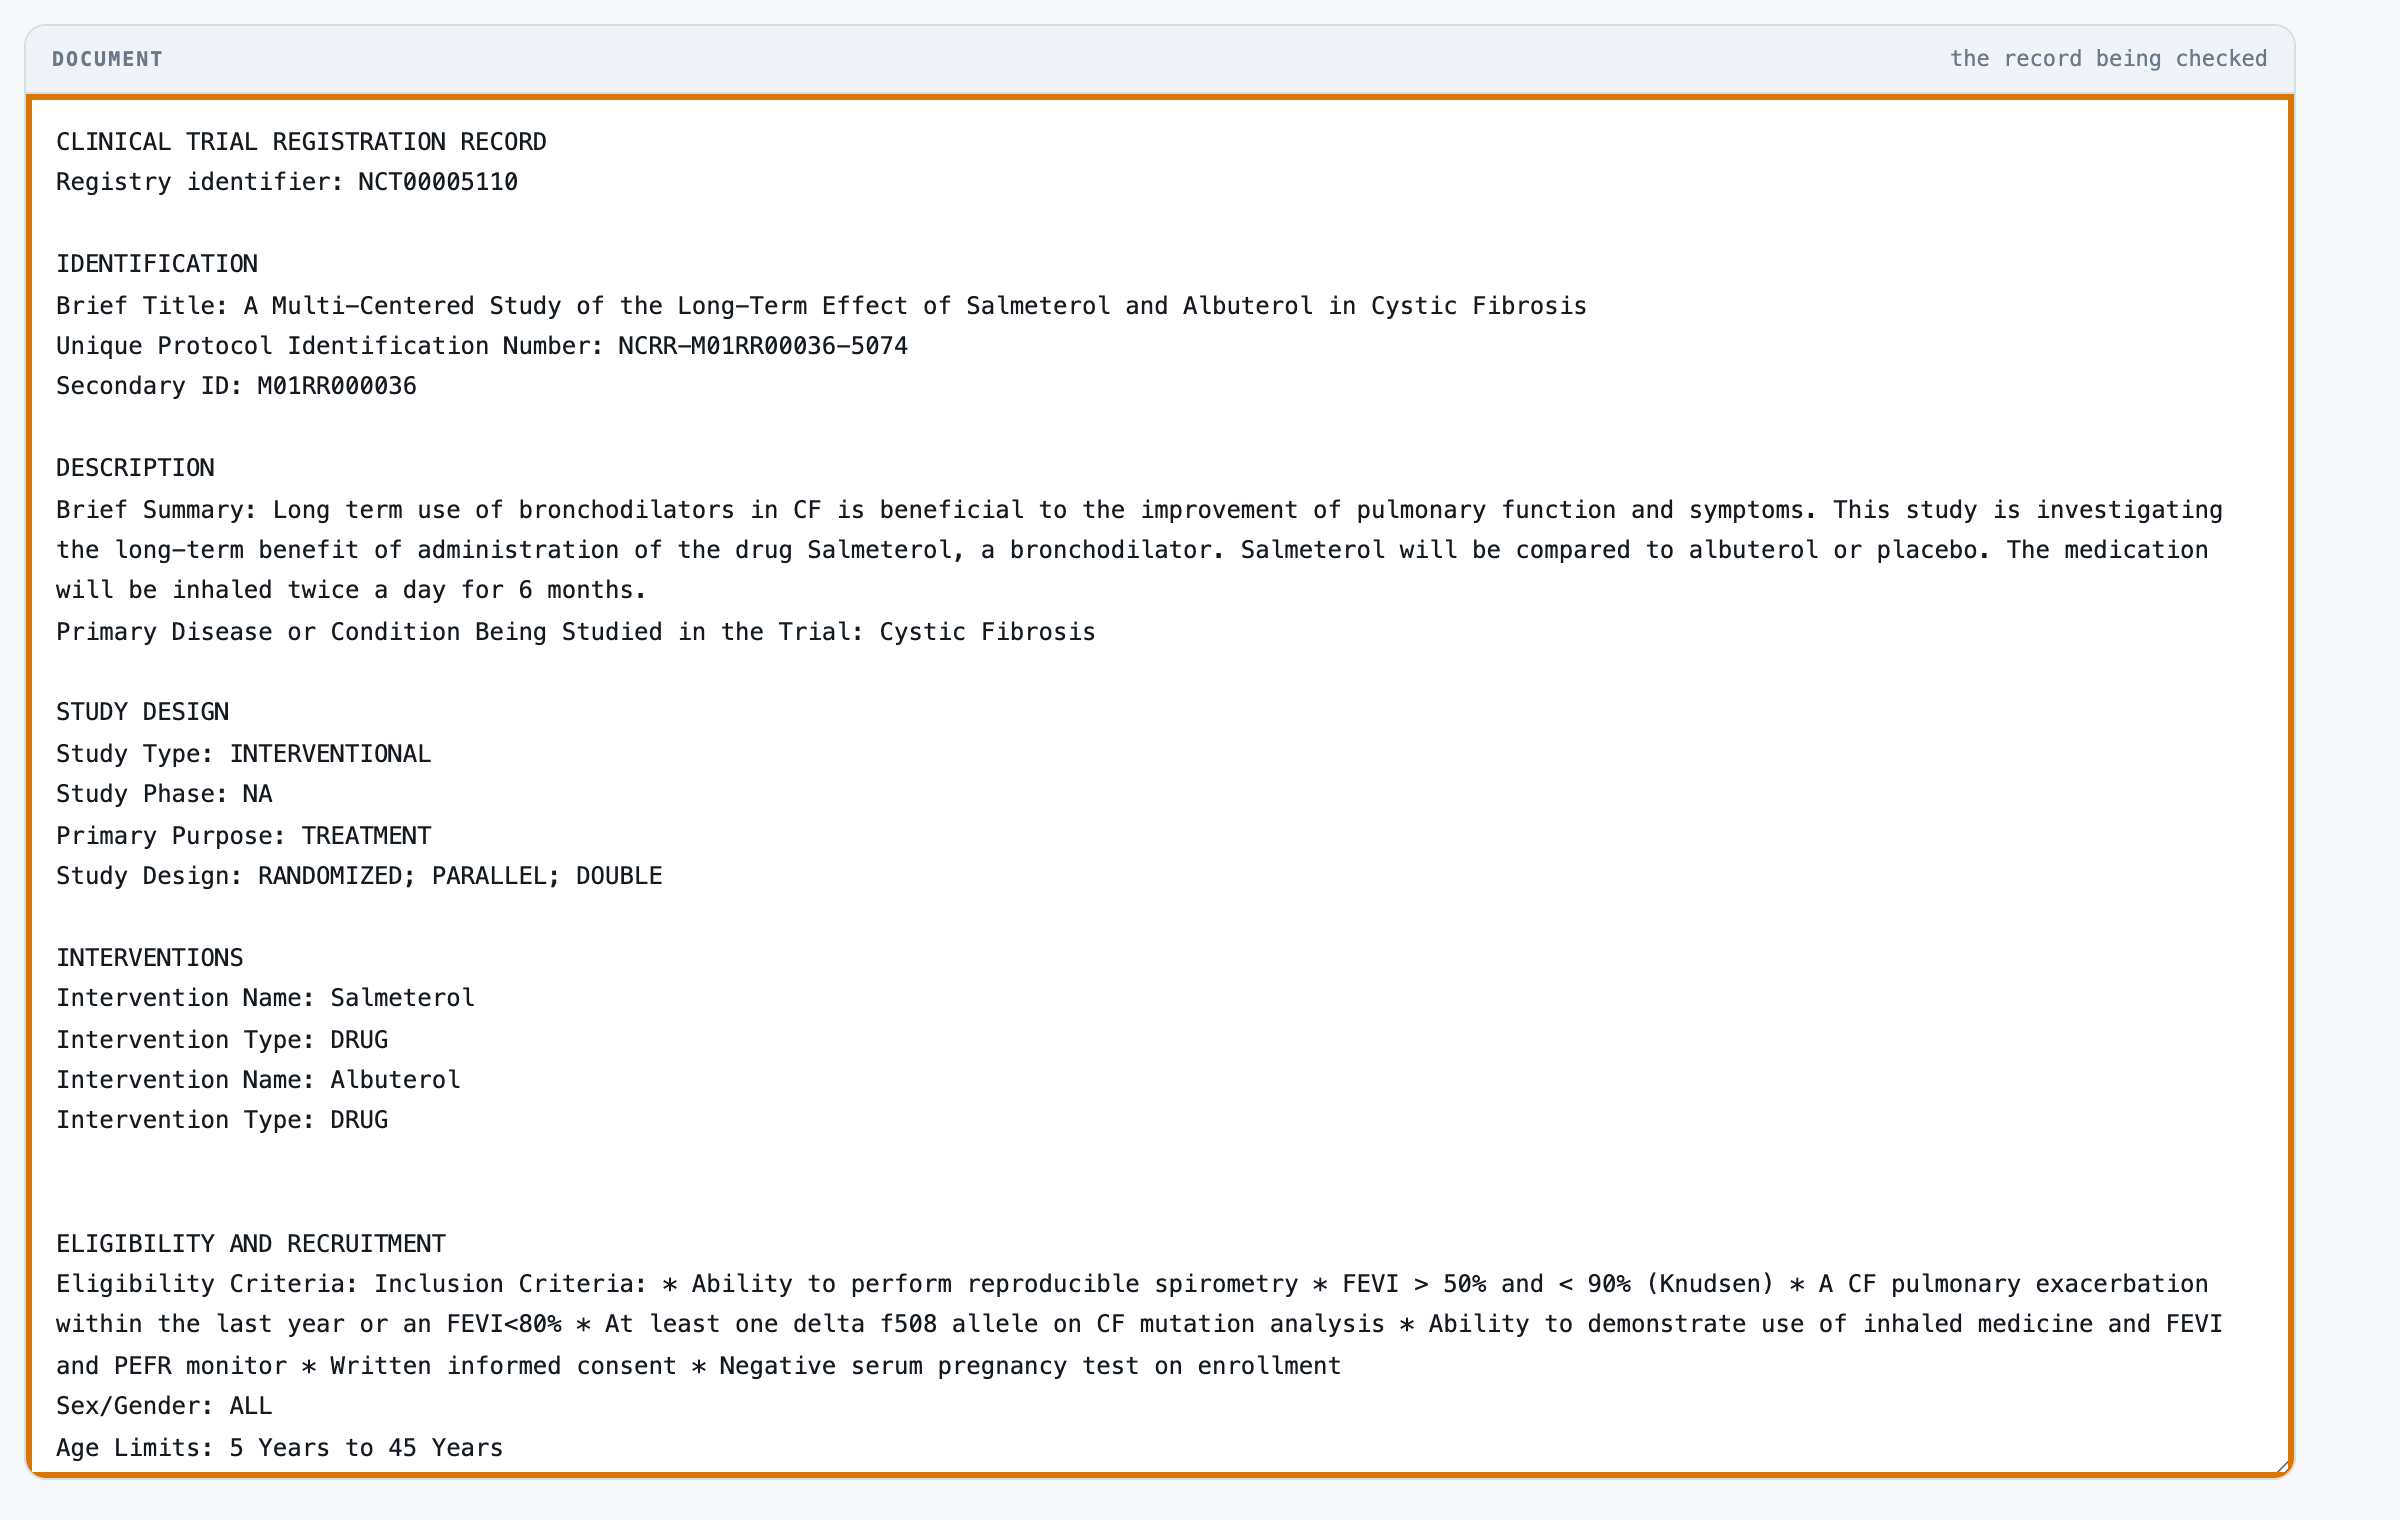
CLINICAL TRIAL REGISTRATION RECORD
Registry identifier: NCT00005110

IDENTIFICATION
Brief Title: A Multi-Centered Study of the Long-Term Effect of Salmeterol and Albuterol in Cystic Fibrosis
Unique Protocol Identification Number: NCRR-M01RR00036-5074
Secondary ID: M01RR000036

DESCRIPTION
Brief Summary: Long term use of bronchodilators in CF is beneficial to the improvement of pulmonary function and symptoms. This study is investigating the long-term benefit of administration of the drug Salmeterol, a bronchodilator. Salmeterol will be compared to albuterol or placebo. The medication will be inhaled twice a day for 6 months.
Primary Disease or Condition Being Studied in the Trial: Cystic Fibrosis

STUDY DESIGN
Study Type: INTERVENTIONAL
Study Phase: NA
Primary Purpose: TREATMENT
Study Design: RANDOMIZED; PARALLEL; DOUBLE

INTERVENTIONS
Intervention Name: Salmeterol
Intervention Type: DRUG
Intervention Name: Albuterol
Intervention Type: DRUG


ELIGIBILITY AND RECRUITMENT
Eligibility Criteria: Inclusion Criteria: * Ability to perform reproducible spirometry * FEVI > 50% and < 90% (Knudsen) * A CF pulmonary exacerbation within the last year or an FEVI<80% * At least one delta f508 allele on CF mutation analysis * Ability to demonstrate use of inhaled medicine and FEVI and PEFR monitor * Written informed consent * Negative serum pregnancy test on enrollment
Sex/Gender: ALL
Age Limits: 5 Years to 45 Years
Accepts Healthy Volunteers: No
Overall Recruitment Status: SUSPENDED

DATES
Record Verification Date: 2001-11

REGULATORY
Availability of Expanded Access: No

SPONSOR, RESPONSIBLE PARTY AND SITES
Name of the Sponsor: National Center for Research Resources (NCRR)
Facility Information: Division of Allergy and Pulmonary Medicine, St Louis, United States